
Fraction of signal in features
- In the app
- Analysis panel, under 1D tasks: Fraction of signal in features
- Command
chromosomix frip- Reads
- Tracks and interval files, with or without a contact map
- Also known as
- FRiP, reads in peaks, enrichment
Fraction of signal in features is FRiP, the fraction of reads in peaks, taken of any loaded track: the share of its signal inside a set of intervals, beside the share of the genome the set covers. Their ratio, the enrichment, is the column to read. Use it to check a library against its peaks, or to rank tracks against a set.
Running it
Section titled “Running it”Load one or more tracks and a feature set (Tracks and features). Open Fraction of signal in features on the 1D tasks card of the Analysis panel, tick the tracks and choose the features; no map is needed. The drawer shows a row per track, the strongest enrichment first.
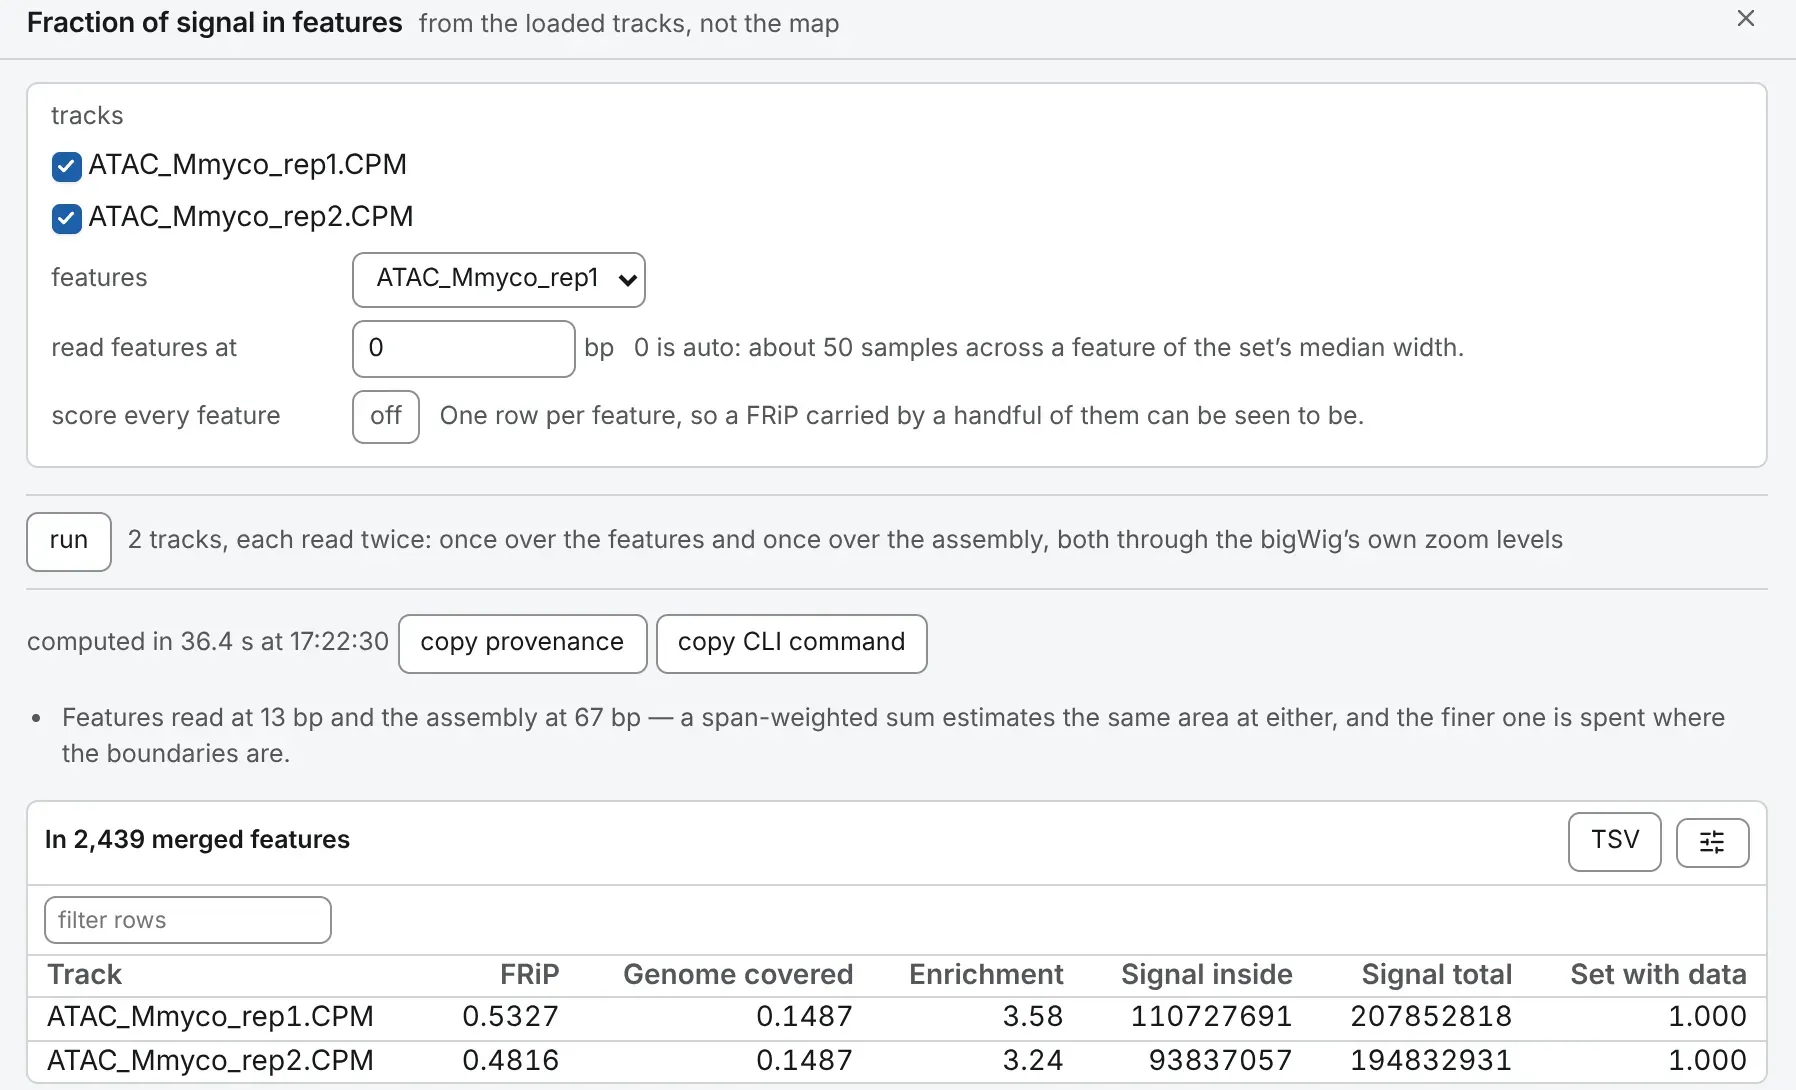
The same run from a shell:
chromosomix frip [<map>] --tracks <tracks.bw> --features <features.bed> [--<parameter> <value> …] --out <dir>Parameters
Section titled “Parameters”read features at 0, the default, reads about 50 samples across a feature of the set’s median width. score every feature adds a table with each feature’s signal, mean and share of the track’s total, which shows whether the FRiP belongs to the set or to a handful of its features.
| Parameter | Takes | Default | What it does |
|---|---|---|---|
tracks--tracks <file,…> | files: a signal track | — | — |
features--features <file> | a file: a feature set | — | — |
read features at--resolution <n> | a whole number, at least 0, in bp | 0 | 0 is auto: about 50 samples across a feature of the set’s median width. |
score every feature--perFeature / --no-perFeature | on or off | off | One row per feature, so a FRiP carried by a handful of them can be seen to be. |
What it writes
Section titled “What it writes”- table: a table: TSV
On the command line every file goes into --out as <file stem>.frip.<output>.<ext>, and each TSV opens with # lines naming the build, the file and every parameter that produced it.
Reading the result
Section titled “Reading the result”- FRiP: the signal inside the set over the signal everywhere. Signal is an area, value × bases, so a wide feature weighs by its width.
- Genome covered: the set’s bases over the assembly’s.
- Enrichment: FRiP over genome covered. 1 is a track with no preference for the set: 30 % of the signal in 30 % of the genome. The same 30 % in half a per cent of the genome is a sixtyfold enrichment.
- Signal inside and Signal total: the two areas.
- Set with data: the share of the set’s bases the track has any value for. A low one means the FRiP is about a small part of the set.
The features are merged first, so a base two of them share, as transcripts of one gene do, counts once; the table’s title gives the number of merged features. The whole track is read from a zoom level where the file has one fine enough, and the note gives both resolutions.
Compared with ENCODE’s FRiP
Section titled “Compared with ENCODE’s FRiP”This is the ENCODE guidelines’ FRiP, of a track’s signal rather than of reads. In the validation, reading the whole track from a zoom level moves a FRiP by less than the last of its four decimals.
Documentation of chromosomix v0.20.0 · built from 40ec1ec · source